01 / UniMolRep
Explore a molecule
- Start with
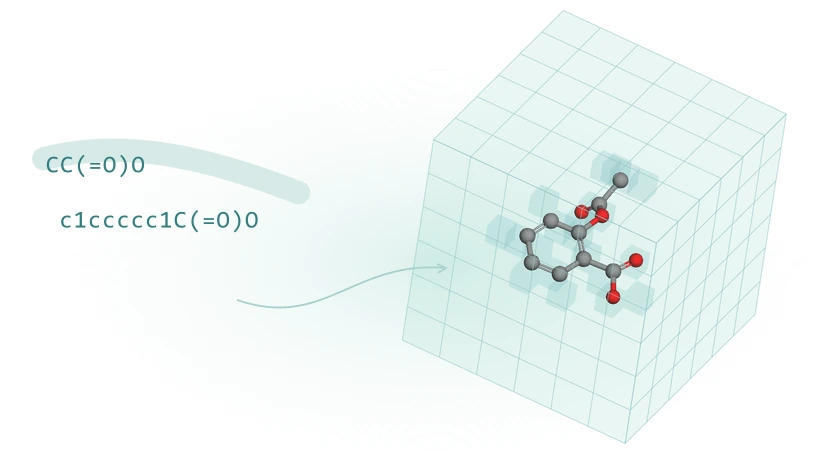
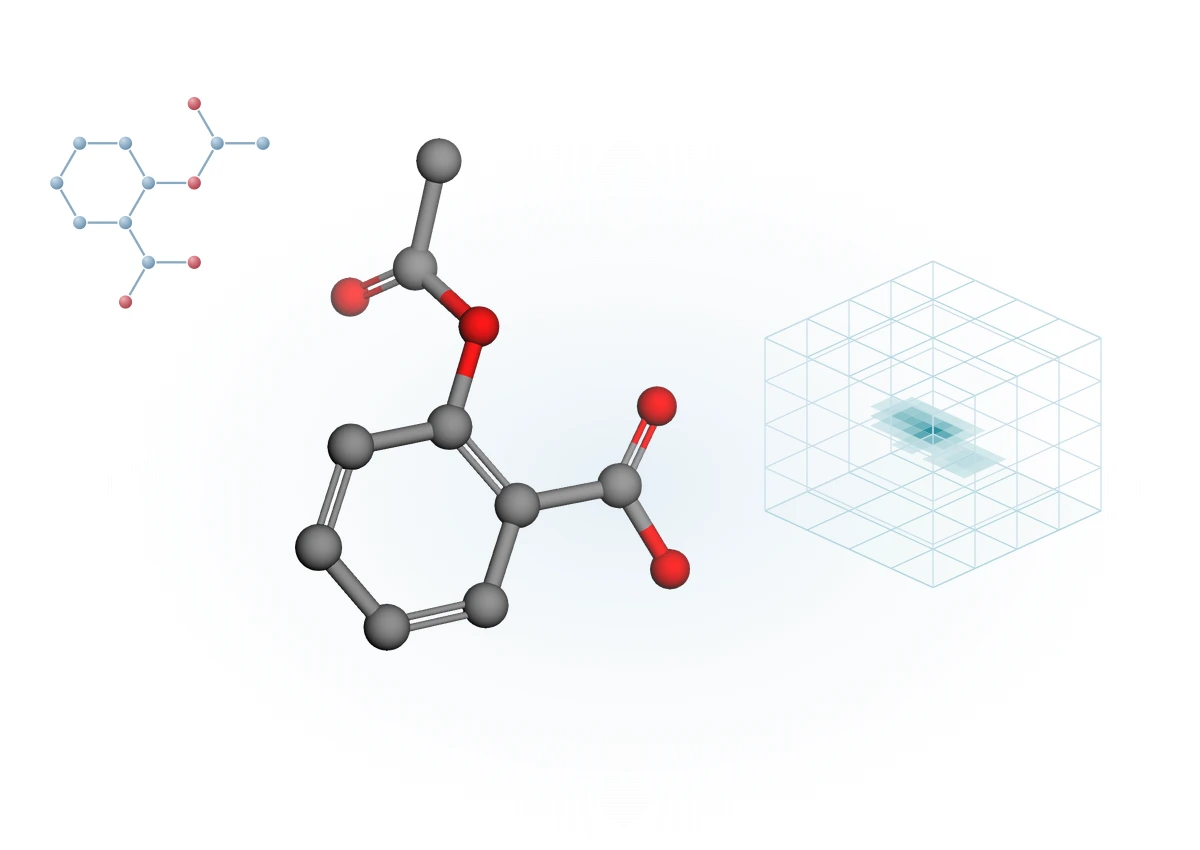
- A molecule as SMILES
- Receive
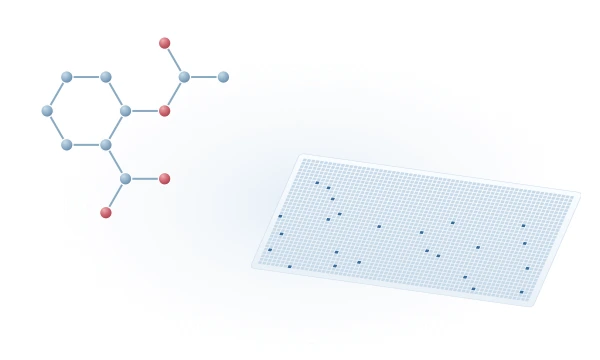
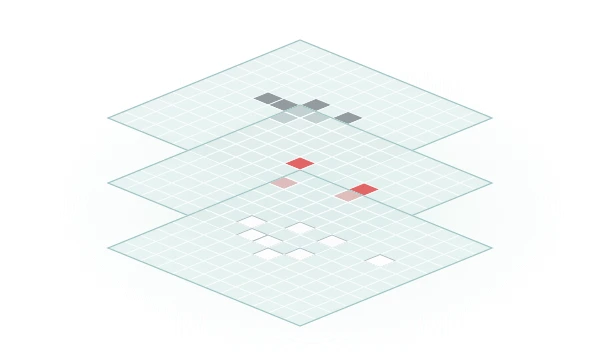
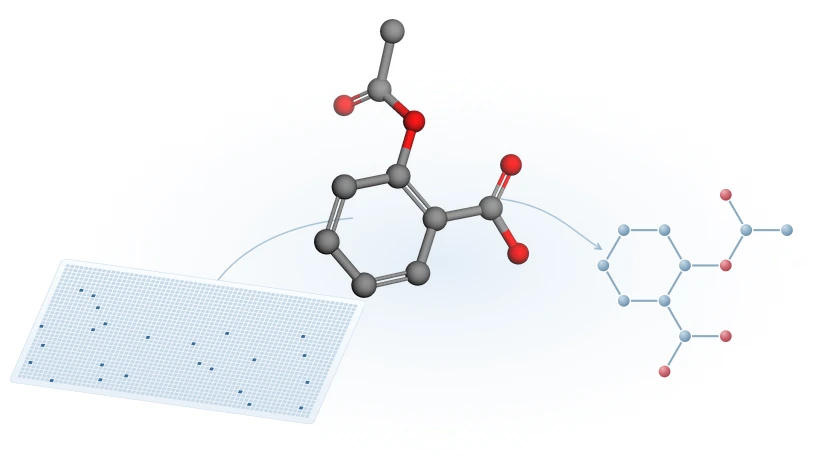
- Fingerprints, atom graphs, geometry and spatial grids
Understand the structure a model sees.
Explore representationsExplore molecular representations, screen against a protein target and score a bound pose using your own inputs. Discover how ETDock predicts binding poses, and connect each method to the research behind it.
Choose your task01 / UniMolRep
Understand the structure a model sees.
Explore representations02 / DL-FSG
Compare molecules within the same target.
Screen a molecule03 / ETDock
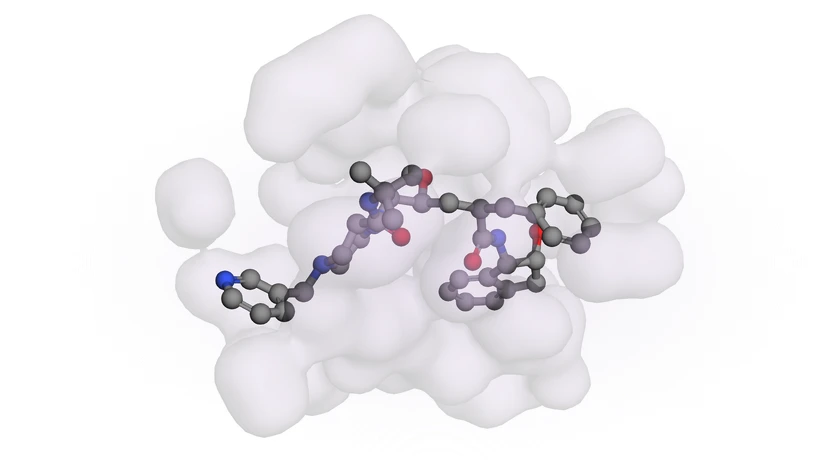
Predict ligand poses in your browser.
Explore docking04 / GNINA / BAP
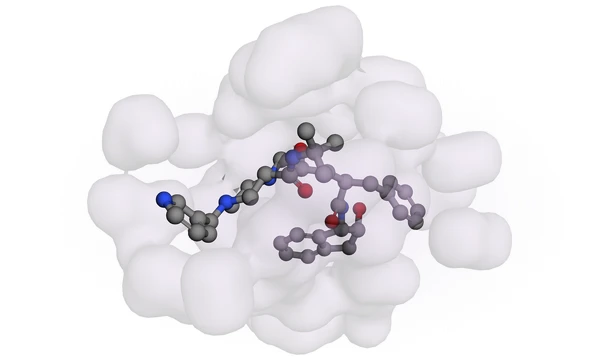
Choose the protein chains and bound ligand.
Score a complexOur research connects molecular representation with three questions: which compounds to prioritise, how they bind, and how strongly they may bind.
Fingerprints, graphs and geometry capture different information. UniMolRep studies how these descriptions support molecular learning.
Learning from sequence and spatial features together helps rank molecules for a protein target. DL-FSG studies this ligand-based screening task.
A candidate molecule’s position and orientation determine how it meets a protein. ETDock learns distances between the partners to reconstruct a binding pose.
Given a protein–ligand complex, AGIMA-Score studies how contacts between the partners can inform an estimate of binding strength.
Research project
The project investigates how molecular grids and deep learning can improve the accuracy, interpretability and efficiency of protein–ligand affinity prediction. Its aim is to support structure-based drug design through better virtual screening and rescoring of protein–ligand poses.
Calculations run in your browser. Molecules and uploaded complexes stay on your device; PDB lookups request the selected structure from the Protein Data Bank.